Using CRISPR as a tool to identify Alzheimer’s Genes and Epigenetic Changes
Why genetic variation is difficult to interpret in Alzheimer’s disease?
Alzheimer’s disease (AD) is one of the most complex neurodegenerative disorders, with its development influenced by a combination of genetic, epigenetic and environmental factors. As the disease progresses, patients suffer from:
- Confusion
- Physical decline
- Social withdrawal
- Depression & Anxiety
- Trouble swallowing
- Severe memory Loss
While genome-wide association studies (GWAS) and epigenome-wide association studies (EWAS) have identified numerous genetic and epigenetic changes associated with Alzheimer’s disease, one fundamental question remains: Do these molecular changes actually cause disease, or are they consequences of the disease process?
Alzheimer’s disease is classified as: Familial and Sporadic forms. Familial Alzheimer’s disease, is a rare, inherited form caused by specific gene mutations, represented by a small proportion of cases. It is most often caused by mutations in genes like APP, PSEN1 and PSEN2. These genes are closely connected with amyloid-beta (Aβ) production and provide relatively clear targets for functional studies.
Sporadic Alzheimer’s disease, on the other hand, is considerably more complicated, that occur in 95% of cases. The sporadic form is influenced by a mix of age, environment, and general risk factors. Common genetic variants individually have relatively small effects, and many Alzheimer’s-associated GWAS variants occur in non-coding regions of the genome. This makes it difficult to determine how a particular variant changes gene regulation or contributes to disease risk.
Epigenetic research adds another layer of complexity. DNA methylation and other epigenetic modifications can alter gene expression without changing the underlying DNA sequence. However, because many Alzheimer’s EWAS studies use post-mortem brain tissue, it can be difficult to determine whether an observed methylation change contributed to Alzheimer’s disease or resulted from ongoing pathology.
This is where gene editing becomes particularly valuable.
Literature research from University of Exeter Medical School, U.K have explored how CRISPR-Cas gene-editing technology could help answer this question. Rather than simply identifying genetic associations, CRISPR allows researchers to experimentally manipulate specific DNA sequences and observe the resulting biological consequences. This creates an opportunity to move from correlation toward functional understanding of Alzheimer’s disease mechanisms.
Using CRISPR as a functional genomics tool
The research brings together evidence from gene-editing studies, Alzheimer’s disease genetic research, epigenomic studies to recognise CRISPR technology including zinc-finger nucleases, TALENs and CRISPR-Cas9, as a tool in genomics.
Traditional CRISPR-Cas9 can introduce a double-strand DNA break at a selected genomic location. The cell can repair this break through non-homologous end joining (NHEJ), which can introduce insertions or deletions and effectively disrupt a gene. Alternatively, homology-directed repair (HDR) can introduce a defined sequence change when a suitable donor template is supplied.
This creates two particularly useful research strategies:
- Gene disruption
- Precise genetic correction.
Using CRISPR’s multiplexing capability, multiple guide RNAs can be used to target several genomic loci, potentially enabling researchers to investigate combinations of Alzheimer’s-associated variants rather than studying each variant in isolation.
CRISPR creates better Alzheimer’s disease models
CRISPR has been recognised for the creation of isogenic cell models.
For this, induced pluripotent stem cells (iPSCs) carrying an Alzheimer’s-associated mutation are selected and these mutations are corrected using CRISPR. The resulting cells can then be compared with the original patient-derived cells.
Because the two cell lines are genetically almost identical apart from the targeted mutation, differences in cellular behaviour can be more confidently attributed to that specific genetic change.
This approach has already produced important findings in familial Alzheimer’s disease.
For example,
- Correction of the PSEN2 N141I mutation in patient-derived iPSC neurons normalized the Aβ42/Aβ40 ratio.
- Gene-corrected isogenic iPSC models have been generated for PSEN1 A79V and L150P mutations.
These models can be used to study disease mechanisms, test potential drug targets and investigate how specific mutations influence neuronal biology, making their practical value substancial.
Targeting gene APP and amyloid-beta production
CRISPR has also been used to investigate the APP–BACE1–amyloid pathway, one of the best-known biological pathways associated with Alzheimer’s disease.
Studies describes the research involving the APP Swedish mutation, a well-characterized amyloidogenic mutation associated with increased Aβ40 and Aβ42 production. In patient-derived fibroblasts carrying the mutation, disruption of the relevant APP cleavage site resulted in an approximately 60% reduction in Aβ production. The same mutation was subsequently targeted in cultured neurons and in an Alzheimer’s mouse model, although editing efficiency in the mouse model was limited by the high copy number of the APP transgene.
Another strategy targeted the C-terminus of APP to interfere with its endosomal trafficking. The resulting experiments showed reduced β-secretase cleavage and decreased Aβ secretion in iPSCs and mouse-derived cells.
These findings demonstrate an important practical principle where CRISPR can be used to investigate specific molecular steps leading to pathological Aβ accumulation, rather than simply targeting an entire gene.
APOE4: Demonstrating cell-specific effects
One of the most informative applications of CRISPR is the involvement of APOE ε4, the strongest common genetic risk factor for sporadic Alzheimer’s disease.
Findings showcase that CRISPR-Cas9-mediated HDR to convert an APOE ε3 iPSC line into a homozygous APOE ε4 line. They then differentiated these iPSCs into different brain cell types, allowing them to investigate the effects of APOE4 in neurons, astrocytes and microglia.
The results revealed striking cell-type-specific effects.
Compared with APOE3 cells, APOE4 astrocytes and microglia demonstrated impaired Aβ clearance, while APOE4 neurons showed increased Aβ42 secretion.
However, converting APOE4 to APOE3 in iPSCs derived from sporadic Alzheimer’s patients resulted in reduced extracellular Aβ uptake and accumulation.
These findings are a powerful demonstration of how gene editing can move genetic epidemiology toward biological explanation.
Instead of simply knowing that APOE4 increases Alzheimer’s risk, researchers can experimentally change the allele and observe how that variant alters cellular behaviour in different brain cell types.
Moving beyond genetic mutations to epigenetics
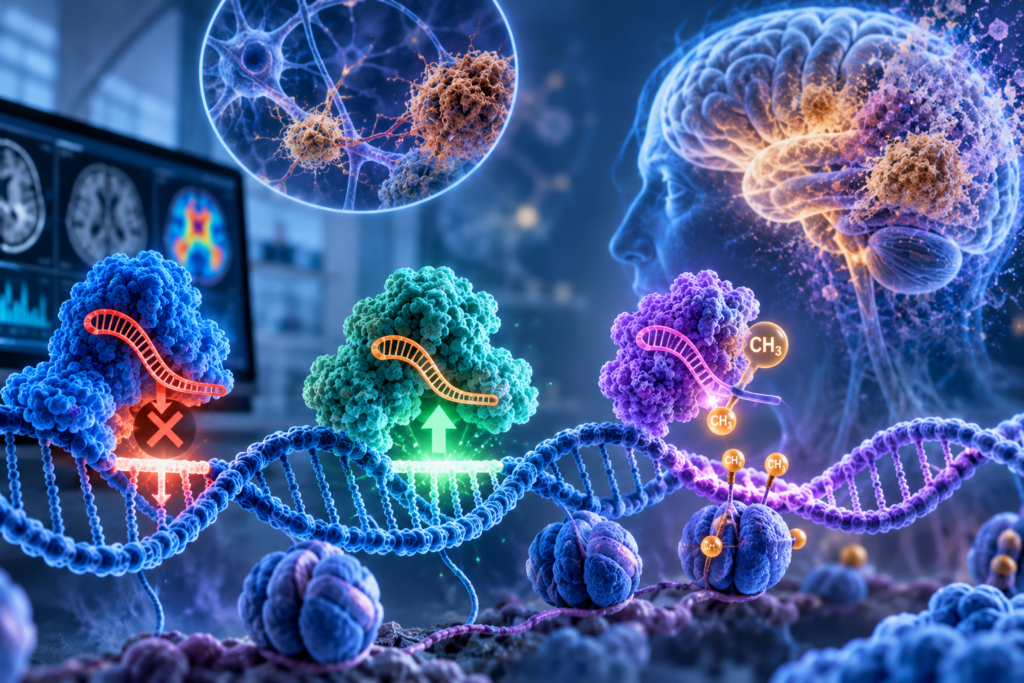
Unlike conventional CRISPR-Cas9, which can permanently alter DNA sequence, modified Cas9 proteins called dCas9 can be engineered to bind specific DNA sequences without cutting them.
These systems can be coupled to molecular effectors that modify gene expression or DNA methylation.
For example, CRISPR interference (CRISPRi) uses dCas9 linked to transcriptional repressors such as KRAB to silence gene expression. Conversely, CRISPR activation (CRISPRa) can increase transcription through activating domains.
DNA-modifying enzymes such as DNMT3A or TET1 can also attached to dCas9 where DNMT3A can add methyl groups to DNA, whereas TET1 can promote DNA demethylation.
These technologies offer an exciting possibility for Alzheimer’s research where experimentally recreating disease-associated epigenetic states and determining whether they produce Alzheimer’s-related cellular phenotypes.
Investigating ANK1 and Alzheimer’s-associated methylation
ANK1 is one the critical gene at an epigenetic locus repeatedly associated with Alzheimer’s disease.
EWAS studies have identified differential methylation at ANK1 and other loci, including CDH23 and RHBDF2. However, association alone cannot establish causality.
CRISPR-based epigenome editing could potentially address this problem by deliberately inducing Alzheimer’s-associated methylation patterns at specific loci in otherwise healthy cells. The changes in gene expression, neuronal function and disease-related cellular characteristics can be further measured.
Supposedly, if introducing a particular methylation pattern produces an Alzheimer’s-associated phenotype, it would provide stronger evidence that the epigenetic alteration may contribute causally to disease. If it produces no meaningful phenotype, the modification could instead represent a consequence or biomarker of disease.
High-throughput CRISPR screening could accelerate discovery
As evident, Alzheimer’s disease is unlikely to be driven by a single genetic or epigenetic change.
CRISPR’s ability to perform high-throughput pooled screens could therefore become particularly important.
Large sgRNA libraries can target thousands of genes or regulatory elements simultaneously. Cells can then be separated according to a measurable phenotype, while sequencing identifies which guide RNAs are enriched in particular cell populations.
Combining CRISPR screening with single-cell RNA sequencing, as demonstrated by approaches such as Perturb-seq, could allow researchers to connect individual genetic perturbations with changes in cellular states and gene-expression networks.
This could be particularly useful for Alzheimer’s disease because different cell types—including neurons, astrocytes and microglia—may respond differently to the same genetic or epigenetic change.
The potential of multiplexed epigenome editing
CRISPR could reproduce multiple Alzheimer’s-associated epigenetic changes simultaneously.
Guide RNA libraries can be designed based on reproducible EWAS findings and introduce selected methylation changes into non-Alzheimer’s cells.
Further, Subsequent transcriptomic and epigenomic analysis could determine whether these alterations collectively produce Alzheimer’s-associated phenotypes. Arrayed CRISPR screening could go even further by placing different guide combinations into individual wells. This could help identify synergistic interactions between genetic and epigenetic loci.
Such approaches could be particularly valuable for sporadic Alzheimer’s disease, where multiple small-effect variants and regulatory changes may interact.
Merits of using CRISPR for Alzheimer’s research
Better disease models: CRISPR-corrected iPSCs can create genetically matched disease and control cell lines.
Variant validation: Cellular behaviour due to Alzheimer’s-associated variants can be determined.
Drug discovery: Genetically defined disease models can be used to identify and test therapeutic targets.
Biomarker research: Functional validation can help distinguish causal molecular signatures from secondary disease changes.
Epigenetic research: dCas9-based tools can experimentally manipulate DNA methylation and gene regulation at specific loci.
High-throughput screening: Pooled CRISPR and single-cell technologies could accelerate the functional annotation of thousands of genetic and regulatory elements.
Personalized disease modelling: Patient-derived iPSCs combined with targeted gene correction offer a route toward studying individual genetic backgrounds.
Some Challenges worth addressing
Despite its potential, CRISPR come with some challenges that needs to be addressed.
1) Off-target editing
Unintended genetic or epigenetic modifications can complicate interpretation and raise safety concerns. Improved guide-design algorithms, alternative Cas9 enzymes and inducible systems may help address these problems.
2) Delivery
Particularly when attempting to manipulate neurons in vivo.
3) Biological
Sporadic Alzheimer’s disease develops over many years and is influenced by aging and environmental factors. Reproducing this gradual process in relatively short-lived cell cultures is difficult.
Furthermore, many Alzheimer’s-associated variants are located in non-coding regions. Understanding their effects may require analysis of chromatin architecture, enhancer-promoter interactions and three-dimensional genome organization, rather than simply examining individual genes.
Future Insights: From genetic association to biological causality
CRISPR could help transform Alzheimer’s disease research from association-based genomics to functional genomics.
Future combinations of CRISPR-Cas editing, CRISPRi/CRISPRa, targeted DNA methylation editing, multiplexed screening, single-cell sequencing and three-dimensional genome analysis could provide increasingly detailed maps of Alzheimer’s disease biology.
These ultimate practical benefit of CRISPR Technology caould substantially identify which molecular changes are truly causal, could reveal new therapeutic targets, and improve disease models and guide the development of more precise interventions.
CRISPR is therefore not only an established cure for Alzheimer’s disease but a powerful experimental platform that may help researchers understand why disease develops and, ultimately, where intervention may be most effective.